Adsorption of Ethylene on Neutral, Anionic and Cationic Gold Clusters
Andrey Lyalin
[email protected]
Tetsuya Taketsugu
[
Abstract
The adsorption of ethylene molecule
on neutral, anionic and cationic gold clusters consisting of up to 10 atoms
has been investigated using density-functional theory.
It is demonstrated that C22+\nolinebreakH42+ can be adsorbed
on small gold clusters in two different configurations,
corresponding to the π- and di-σ-bonded species.
Adsorption in the π-bonded mode dominates over the di-σ
mode over all considered cluster sizes n, with the exception of the
neutral C22+\nolinebreakH42+−\nolinebreakAu52+ system.
A striking difference is found in the size-dependence of the
adsorption energy of C22+\nolinebreakH42+ bonded to the neutral gold clusters in the
π and di-σ configurations.
The important role of the electronic shell effects in the di-σ mode
of ethylene adsorption on neutral gold clusters is demonstrated.
It is shown that the interaction of C22+\nolinebreakH42+ with small gold clusters
strongly depends on their charge.
The typical shift in the vibrational frequencies of C22+\nolinebreakH42+
adsorbed in the π- and the di-σ configurations
gives a guidance to experimentally distinguish between the two modes of adsorption.
\altaffiliation
On leave from: Institute of Physics, St Petersburg State University,
198504 St Petersburg, Petrodvorez, Russia
Hokkaido University]Division of Chemistry, Graduate School of Science,
Hokkaido University, Sapporo 060-0810, Japan
1 Introduction
The adsorption of unsaturated hydrocarbons on transition metal surfaces has been
studied extensively in order to understand the nature of
hydrocarbon – metal interaction and chemical processes
on solid surfaces; see, e.g., Refs.
1, 2, 3, 4, 5, 6 and
references therein.
The most significant attention was paid to
investigation of ethylene adsorption,
7, 11, 12, 13, 14, 15, 10, 16, 8, 17, 9
because it is the simplest alkene containing an isolated carbon–carbon
double bond. Hence it can be treated as a prototype to study the interaction and reactivity
of different alkenes on metal surfaces. Moreover, the ethylene epoxidation is one of
the most important processes in the chemical industry, because the product of such a reaction –
ethylene oxide is widely used in various applications;
see, e.g., Refs. 18, 19, 20, 21, 1, 2
and references therein.
Ethylene can adsorb on metal surfaces in two different
configurations.11, 12, 14, 1
The first one is the π mode where one metal atom on the surface
is involved in the adsorption of ethylene via a π-bonding.
The second one is the di-σ-bonded mode, when two metal atoms are involved
in the adsorption via a σ bonding.
It was found that the di-σ mode of adsorption is
characterized by the increasingly important role of sp3
hybridization in ethylene, while sp2 hybridization (typical for free molecules)
remains unchanged for π-bonded species.11, 12, 14
The bonding of ethylene with transition metals involves electron transfer from
the filled bonding π orbital of the ethylene to the
metal, alongside a back-donation from the d-orbital of transition metal to the
empty π∗ anti-bonding orbital of ethylene in accordance with the
Dewar-Chatt-Duncanson model 22.
Thus the appearance of the sp3 character of hybridization in
the di-σ-bonded ethylene can be explained by
the increasing role of the electron back-donation from the metal to the
π∗ anti-bonding orbital of ethylene.
Hence the di-σ-bonded ethylene is activated more strongly in comparison with
the π-bonded one, thus it can be more reactive.11, 12
Therefore, an understanding of the specific mechanisms of ethylene adsorption on metal
surfaces is important in order to gain a better insight of the catalytic processes and
reactivity of adsorbed hydrocarbons.
Surprisingly, not many works have been devoted to the investigation
of ethylene adsorption on metal clusters and nanoparticles,
despite the fact that chemical and physical properties of matter
at nanoscale are very different from those of the corresponding
bulk solids. These properties are often controlled by quantum size effects and
significantly depend on the size and structure of atomic clusters.23
A remarkable example is gold.
It is well known that gold in its bulk form is a catalytically inactive and inert metal.
However gold at nanoscale manifests extraordinary catalytic activity
which increases with a decrease in the cluster size of up to 1-5 nm
24, 25, 26, 27.
Moreover, recent studies demonstrate that catalytic activity of gold clusters adsorbed on
an iron oxide support correlates with the presence
of very small clusters of ∼ 10 atoms 28.
It was also reported that gold clusters with the number of atoms
n=3−11 possess extraordinarily high electrocatalytic activity toward the O22+
reduction reaction in acid solutions.29
The origin of such size-dependent catalytic activity of gold remains highly
debated and has yet to be fully understood.
A comprehensive survey of the field can be found in review papers and books;
see, e.g., Refs. 23, 30, 31, 32, 33, 34.
It was demonstrated that the catalytic activity of gold
clusters depends on the type of the support material, the
presence of defects in the material (e.g. F-center defects),
the environment or special additives;
see, e.g., Refs. 25, 35, 36, 31, 37
and references therein.
However, experiments indicate that gold clusters
deposited on an inert support
can efficiently catalyze the process of styrene oxidation
by dioxygen 27. Hence, the origin of catalytic activity derives from the
cluster itself and therefore even free clusters can be effective catalysts.
Currently, a large variety of catalytic reactions on gold clusters
has been studied experimentally. This includes the processes of catalytic oxidation
of carbon monoxide, as well as more complex reactions such as alcohol oxidation,
the direct synthesis of hydrogen peroxide and alkene
epoxidation.27, 38, 26, 39
However, theoretical studies of the gold nanocatalysis
have mainly focused on the investigation of adsorption and catalytic reaction of
O22+ and CO.31, 36
Nonetheless, the theoretical study of the interaction of alkenes with
gold clusters has been relatively unexplored
with the exception of works on the binding of propene on gold and
mixed gold-silver clusters40, 41,
propene epoxidation42 and our recent work
on cooperative adsorption of oxygen and ethylene on small
gold clusters43.
In this paper we report the results of a systematic theoretical study
of the adsorption of C22+\nolinebreakH42+ on free neutral, anionic and cationic
gold clusters consisting of up to 10 atoms.
We find that C22+\nolinebreakH42+ adsorbs on small gold clusters in two different configurations,
corresponding to the π- and di-σ-bonded species.
Adsorption in the π-bonded mode dominates over di-σ
mode over all considered cluster sizes n with the exception of the
neutral C22+\nolinebreakH42+−\nolinebreakAu52+ system, where the
di-σ-bonded configuration is energetically more favorable.
We find a striking difference in the size dependence of the
adsorption energy of C22+\nolinebreakH42+ bonded to neutral gold clusters in the
π and di-σ configurations.
Thus, the adsorption energy of the π-bonded C22+\nolinebreakH42+ develops
non-monotonically as a function of cluster size with a local minima at n=6.
The adsorption energy, calculated for the di-σ-bonded C22+\nolinebreakH42+ exhibits
pronounced odd-even oscillations, showing the importance of the electronic
shell effects in the di-σ mode of ethylene adsorption on gold clusters.
We also demonstrate that the interaction of C22+\nolinebreakH42+ with small gold clusters
strongly depends on their charge. The excess of a positive or a negative charge on
the gold cluster can change the balance between electron donation and
back-donation processes.
Hence ethylene adsorption and reactivity
can be manipulated by the cluster charge.
This effect can be particularly important for understanding the mechanisms of catalytic
activation of ethylene adsorbed on small gold clusters.
2 Theoretical Methods
The calculations are carried out using density-functional theory (DFT).
The hybrid Becke-type three-parameter
exchange density functional paired with the gradient-corrected
Perdew-Wang 91 correlation functional (B3PW91) 44 is used.
The choice of the B3PW91 functional is consistent with our previous work
on cooperative adsorption and catalytic oxidation of ethylene on
small gold clusters.43
The obtained theoretical data on dissociation energies and bonding in
Au22+ (1.904 eV, 2.546 Å), and
C22+\nolinebreakH42+ (7.596 eV, 1.324 Å) are in a good agreement
with those of earlier experimental studies,
Au22+ (2.30 eV, 2.472 Å) 45, and
C22+\nolinebreakH42+ (7.76 eV, 1.339 Å) 46.
The standard LANL2DZ basis set of primitive Gaussians is used to
expand the gold cluster orbitals formed by the 5s25p65d106s1
outer electrons of Au (19 electrons per atom).
The remaining 60 core electrons of the Au atom are represented
by the Hay-Wadt effective core potential accounting for relativistic effects
47. For carbon and hydrogen, the aug-cc-pVTZ basis set48 is employed.
Calculations have been carried out with the use of
the Gaussian 03 code 49.
The structural properties of neutral, anionic and cationic gold clusters have been the subject
of numerous theoretical investigations; see, e.g., Refs. 30, 31, 32
for a review.
In the present work the cluster geometries have been determined with the use of the
cluster fusion algorithm50 which belongs to the class
of genetic global optimization methods 51, 52.
We have successfully used a similar
approach to find the optimized geometries of various types of atomic clusters
53, 54, 55.
The optimized structures of the gold clusters Aun2+, {{\text{Au}_{\vphantom{\text{}}\text{n}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}}, and Aun2++
(n=1−10) obtained in the present work
are presented in 1. These structures
are in a good agreement with those reported in previous theoretical studies; see, e.g.,
Refs. 56, 57, 58, 59, 32.
In order to obtain the most stable configuration of C22+\nolinebreakH42+
adsorbed on neutral, anionic and cationic gold clusters,
we have created a large number of starting geometries by adding C22+\nolinebreakH42+ molecule
in different positions (up to 30) on the surface of the most stable cluster
and up to eight isomer structures of the corresponding Aun2+, {{\text{Au}_{\vphantom{\text{}}\text{n}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}}, and Aun2++
clusters. The starting structures have been optimized further without any geometry constraints.
3 Theoretical results
3.1 Geometry optimization for ethylene adsorbed on neutral, cationic and anionic
gold clusters
The results of the cluster geometry optimization
for neutral C22+\nolinebreakH42+−\nolinebreakAun2+,
anionic {{\text{C}_{\vphantom{\text{}}\text{2}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{}}\nolinebreak\text{H}_{\vphantom{\text{}}\text{4}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{}}{-}\text{}\nolinebreak\text{Au}_{\vphantom{\text{}}\text{n}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}}
and cationic C22+\nolinebreakH42+−\nolinebreakAun2++ clusters
within the size range n≤10
are shown in 2, 3 and 4,
respectively.
Gold clusters with adsorbed ethylene possess various isomer forms whose number
grows dramatically with cluster size.
Among the huge variety of different isomers, one can distinguish
two specific adsorption modes that correspond to the
π- and di-σ configurations of the adsorbate.
The appearance of two different modes of ethylene adsorption on small gold clusters is
similar to that reported for ethylene adsorption on Au(100) and Au(111) surfaces14.
However in the case of adsorption on the bulk, there
are only two possible geometrical configurations of ethylene (we do not consider here
μ-bridging species14,
because ethylene adsorption in such a configuration is not possible
in the case of small clusters). Moreover, there is no structural changes
of the bulk surface due to adsorption of ethylene.
In the case of ethylene adsorption on small gold clusters,
each of the π- and di-σ configurations possess a variety of structural isomers.
These isomers correspond to the adsorption in nonequivalent
positions on the surface of clusters.
In 2, 3 and 4
we present the most stable structures of π- and di-σ-bonded
ethylene adsorbed on small gold clusters.
The C – C and C – Au interatomic distances are given in angstroms.
The interaction of gold clusters with ethylene
results in an electron donation from the C22+\nolinebreakH42+
to the gold cluster alongside a back-donation from
the cluster to the adsorbate in accordance with the
Dewar-Chatt-Duncanson model 22.
The rearrangement of the electronic structure and
electron transfers upon adsorption lead to a dramatic
change in cluster geometry. The interplay of electronic
and geometry shell effects, also influence the cluster structure 55.
Free neutral gold clusters possess planar, two-dimensional (2D)
structures when the cluster size is up to 13 Au atoms.60, 59
In 1, we present the most stable structures obtained for Aun,
Aun2++ and {{\text{Au}_{\vphantom{\text{}}\text{n}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}} clusters (n = 1, …, 10)
within the B3PW91/LANL2DZ DFT method.
2 demonstrates that adsorption of C22+\nolinebreakH42+
in the π configuration leads to a prevalence of three-dimensional (3D) structures
of gold clusters for n≥9; the structures of small Au32+ and Au42+ clusters
transform to that which are energetically favorable for free cations
Au32++ and Au42++, respectively.
For n<9, ethylene favorably adsorbs at the cluster edge
where the carbon-carbon bond lay in the plane of the cluster with the exception of the
C22+\nolinebreakH42+−\nolinebreakAu62+ system, where the carbon-carbon bond is located perpendicular
to the cluster plane (see 2).
Adsorption of C22+\nolinebreakH42+ in the di-σ configuration results in a considerable
rearrangement in the structure of neutral gold clusters.
Thus, the geometry of Au32+, and Au62+ – Au102+ clusters changes upon
C22+\nolinebreakH42+ adsorption. It is known that gold clusters possess a large number of
energetically close-lying structural isomers.30, 32
Interaction with adsorbate can easily alter their energy order.
It is important to note, that interaction with adsorbate can be responsible for
structural rearrangements even for the relatively large clusters. Thus,
recent DFT calculations demonstrate that the morphology of Au792+ nanoparticle
can be transformed by interactions with a CO atmosphere.61
It is not surprising that the structures of positively and negatively charged
ionic gold clusters change upon ethylene adsorption.
3 demonstrates that although anionic gold
clusters {{\text{Au}_{\vphantom{\text{}}\text{n}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}} with n = 1, …, 10 remain planar upon
ethylene adsorption (both for the π and di-σ adsorption configurations),
they manifest considerable structural rearrangements.
Thus, gold cluster anions alter their structure upon ethylene adsorption
in the π-configuration for n = 4, 7, 8, and 10.
In the case of di-σ mode of ethylene adsorption,
structural rearrangements occur for gold cluster anions with n = 3 – 8.
A carbon-carbon bond of the adsorbed ethylene lie in a cluster plane
for all structures presented in 3.
4 shows that in the case of ethylene adsorption
on gold cluster cations, Aun2++, the resulting cluster systems
C22+\nolinebreakH42+−\nolinebreakAun2++ exhibit 3D structures for
n= 5 – 10 in the case of the π mode of ethylene adsorption; and n = 8 and 10 in the case
of the di-σ mode. In both cases of the π and di-σ modes of ethylene adsorption,
the structural rearrangements in gold cluster cations occur for n≥ 5.
Such a severe change in the morphology of gold clusters due to the interaction with ethylene
makes the procedure of structural optimization rather complicated.
In order to get energetically favorable structures of
C22+\nolinebreakH42+−\nolinebreakAun2+, {{\text{C}_{\vphantom{\text{}}\text{2}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{}}\nolinebreak\text{H}_{\vphantom{\text{}}\text{4}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{}}{-}\text{}\nolinebreak\text{Au}_{\vphantom{\text{}}\text{n}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}}, and C22+\nolinebreakH42+−\nolinebreakAun2++ clusters, it is
necessary to consider adsorption of ethylene not only
on the most stable structures of the corresponding
Aun2+, {{\text{Au}_{\vphantom{\text{}}\text{n}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}}, and Aun2++ clusters, but also on a large
number of their energetically less favorable isomer states.
In the present work we have taken into account up to
8 isomer structures of the host gold clusters.
This insures that we do not miss energetically favorable structures with considerable
change in cluster geometry due to C22+\nolinebreakH42+ adsorption.
3.2 Energetics of ethylene adsorption on neutral, anionic and cationic gold clusters
We first study the energetics of C22+\nolinebreakH42+ adsorption on neutral gold clusters
Aun with 1≤n≤10.
The spin states of the optimized C22+\nolinebreakH42+−\nolinebreakAun2+ structures
are doublet and singlet for odd and even n, respectively.
5 shows the evolution of the
molecular adsorption energy, Eads, calculated for the most bounded
π and di-σ configurations of adsorbed C22+\nolinebreakH42+
as a function of cluster size n.
The adsorption energy is defined as
[TABLE]
where Etot(M) denotes the total energy of the most stable structure
of the molecule (cluster) "M".
In our previous work, we have studied adsorption of ethylene on small neutral gold clusters
in the π-bonded configuration.43
Here, we demonstrate that C22+\nolinebreakH42+
can be adsorbed on small gold clusters in two different configurations,
corresponding to the π- and di-σ-bonded species.
5 shows that the π-bonded mode of C22+\nolinebreakH42+ adsorption is energetically favorable
in the considered range of cluster sizes n, with the
exception of a single Au atom
which interacts with one carbon atom in C22+\nolinebreakH42+, forming a σ bond,
and the Au52+ cluster, where the di-σ-bonded configuration of
C22+\nolinebreakH42+−\nolinebreakAun2+ is 0.05 eV more bounded compared with the π-bonded one.
It is seen from 5 that the adsorption energy of
the π-bonded C22+\nolinebreakH42+ on Aun2+ clusters exhibit a maximum
at n= 3 and 4 followed by a fast drop with a minimum at n= 6.
For 6≤n≤10 the adsorption energy of π-bonded C22+\nolinebreakH42+
shows a weak oscillatory behavior as a function of n.
The n-evolution of the adsorption energy calculated for
the di-σ-bonded C22+\nolinebreakH42+ is considerably different
from that of the π-bonded C22+\nolinebreakH42+.
5 demonstrates that the adsorption energy of
the di-σ-bonded C22+\nolinebreakH42+ has an odd-even oscillatory behavior
with a local maxima for odd numbers of Au atoms n= 3, 5, 7, and 9.
For clusters with an even number of n, the adsorption energy Eads
of the di-σ-bonded C22+\nolinebreakH42+ is small.
Hence, C22+\nolinebreakH42+ readily adsorbs in the di-σ configuration—but
only for neutral gold clusters with an odd number of Au atoms.
It is interesting to note that in the case of C22+\nolinebreakH42+ adsorption on a Au(111) surface,
the di-σ mode (Eadsdi−σ = 0.6502 eV) is
energetically more favorable in comparison to the π mode
(Eadsπ = 0.1545 eV) 14.
A similar effect is observed for C22+\nolinebreakH42+ adsorbed on a Au(100) surface where
theoretical values of Eadsdi−σ = 0.7790 eV and
Eadsπ = 0.2456 eV have been reported 14.
It is known that the type of C22+\nolinebreakH42+ bonding depends upon the surface structure.
It has been found that the π-bonded mode dominates in the adsorption of
low coordinated atoms and step sites 62.
In the case of small metal clusters, almost all atoms are essentially
surface atoms that are undercoordinated as compared to the bulk.63
Thus, the π-bonded configuration of the adsorbed C22+\nolinebreakH42+ dominates for
small gold clusters. As the number of low coordinated atoms
decreases with increase in cluster size, we suppose that
the di-σ mode will become dominant for large gold clusters.
6 and 7
present the n-evolution of the the C–C bond distance, rC−C,
and the total charge transfer to the adsorbate, ΔqC2H4,
obtained from the natural bond orbitals (NBO) population analysis64.
6 shows that the C–C bond in the π-bonded C22+\nolinebreakH42+
is enlarged up to 1.37–1.39 Å in comparison with the case of a
free C22+\nolinebreakH42+ (1.32 Å). The rC−C calculated for the π-bonded C22+\nolinebreakH42+
depends slightly on the cluster size, evolving with n
similar to Eads.
On the other hand, for the di-σ-bonded ethylene the C–C bond
is enlarged up to 1.42–1.50 Å, which is comparable with the length
of a single C–C bond in ethane, C22+\nolinebreakH62+.
The n-dependence of rC−C for the di-σ-bonded C22+\nolinebreakH42+ manifests an irregular
behavior with maxima at n= 2, 4, 7 and 9.
An increase in the C–C bond length, as well as as noticeable change of the
bending of the H atoms for di-σ-bonded C22+\nolinebreakH42+ (see 2),
can be explained by the increasingly important role of the sp3 hybridization
due to the electron donation from the gold cluster to the
anti-bonding π∗ orbital of C22+\nolinebreakH42+. Therefore, rC−C in the di-σ-bonded
ethylene approaches its typical values for a single C–C bond in sp3 hybridized C22+\nolinebreakH62+.
Hence the di-σ-bonded ethylene is activated strongly in comparison with
the π-bonded one, thus it can be more reactive in catalytic processes.
The total charge transfer to the adsorbed C22+\nolinebreakH42+
depends on the balance between the donation and back-donation processes.
7 demonstrates that in the case of
the π mode of adsorption the total charge transfer
ΔqC2H4 is relatively small and
evolves with n similar to rC−C. For the di-σ mode of
adsorption the back-donation from the gold to the
π∗ anti-bonding orbital of ethylene dominates over electron transfer
from C22+\nolinebreakH42+ to the cluster. The total charge transfer
to C22+\nolinebreakH42+ adsorbed in a di-σ configuration is large and varies
in the range of 0.29 - 0.43 a.u. for n= 2 - 9.
It is seen from
7 that ΔqC2H4 calculated
for the di-σ-bonded C22+\nolinebreakH42+
exhibits a pronounced odd-even oscillations as a function of cluster size n with the
exception of n=1. Note, that for n=1 the charge transfer ΔqC2H4
is small because in this case only one Au−C σ bond is formed.
Similar odd-even oscillations have been found in the n-dependence
of the adsorption energy of O22+ on Aun2+ clusters; see, e.g.
56, 65, 23 and references therein.
However, the adsorption energy of CO does not present any
odd-even effects65.
The appearance of the odd-even oscillations in Eads
and ΔqC2H4
for the di-σ mode of adsorption is a result of
the electronic shell effects67, 66, 32
in the gold clusters and can be described within the jellium model.
In spite of its simplicity, the jellium model can explain,
at least on the qualitative level, many physical properties and chemical reactivity of
metal clusters.32, 68, 69
Indeed, each Au atom has one 6s electron which
delocalizes in the whole volume of the cluster.
The delocalized electrons are moving in the
field of a uniform positive charge background, which binds the valence electron cloud.
The gold clusters with an even number of atoms (valence electrons)
have closed electronic shell structure.
Hence these clusters are more stable and less reactive in comparison with
open shell clusters possessing an odd number of atoms (valence electrons).
The prevalence of the electron transfer from the
cluster to the adsorbate (back-donation process) is responsible for the promotion of the
di-σ-mode of C22+\nolinebreakH42+ adsorption.
Indeed, the di-σ mode of C22+\nolinebreakH42+ adsorption on small gold clusters with an
odd number of atoms leads to the transfer of unpaired electron from the gold cluster to the
ethylene anti-bonding π∗ orbital, causing the weakening of the C–C bond.
The neutral gold clusters
with even number of valence electrons possess a closed electronic shell structure,
hence electron transfer to the adsorbate and
formation of the di-σ bonded species is suppressed.
8 demonstrates the evolution of
molecular adsorption energy, Eads and C–C bond distances, rC−C,
as a function of cluster size n for C22+\nolinebreakH42+ adsorbed
on the anionic {{\text{Au}_{\vphantom{\text{}}\text{n}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}} (left column) and cationic Aun2++ (right column)
clusters with 1≤n≤10.
The adsorption energy of C22+\nolinebreakH42+ on a gold cluster depends on the
balance between donation and back-donations processes, hence
it can be manipulated by the cluster charge.
It is seen form 8a and 8b
that excess of the negative or the positive charges on gold clusters
strongly influence the molecular adsorption.
Thus, the interaction of C22+\nolinebreakH42+ with the negatively charged gold clusters {{\text{Au}_{\vphantom{\text{}}\text{n}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}}
becomes weaker if compared with the neutrals;
while excess of the positive charge results in strengthening of the C22+\nolinebreakH42+ and Aun2++ bond.
8a demonstrates that the π-mode of C22+\nolinebreakH42+ adsorption is
not stable for {{\text{Au}_{\vphantom{\text{}}\text{}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}} and {{\text{Au}_{\vphantom{\text{}}\text{2}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}}.
The adsorption energy of the π-bonded C22+\nolinebreakH42+ on the gold cluster anions
{{\text{Au}_{\vphantom{\text{}}\text{n}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}} is 0.17 eV in the case of a trimer {{\text{Au}_{\vphantom{\text{}}\text{3}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}},
and this increases with an increase in the cluster size: up to 0.82 eV at n=7 and dropping
to 0.54 eV at n=8, while slowly increasing with an increase in the cluster size
up to n=10.
8b shows that the size dependence of Eads for
π-bonded C22+\nolinebreakH42+ on the gold cluster cations Aun2++
is very different from that of {{\text{Au}_{\vphantom{\text{}}\text{n}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}}.
Thus, C22+\nolinebreakH42+ readily adsorbs on the cationic
monomer Au2++ and dimer Au22++ with an adsorption energy of
2.95 eV and 3.58 eV, respectively. Further increase in cluster size results in
the sharp decrease in Eads up to 2.06 eV for n=3 followed by
its slow decrease for 3≤n≤6. At n=7 the adsorption energy Eads
exhibits a maximum followed by the second sharp decrease at n= 8 and 9.
For any n in the range 1≤n≤10, the adsorption energy of
the π-bonded C22+\nolinebreakH42+ on
cationic Eadsπ(Aun+),
neutral, Eadsπ(Aun),
and
anionic, Eadsπ(Aun−),
gold clusters satisfies the relation:
Eadsπ(Aun+)>Eadsπ(Aun)>Eadsπ(Aun−).
Indeed, the excess of the positive charge
on the cluster favours the process of electron transfer from the C22+\nolinebreakH42+
to the gold cluster, which stabilizes the π mode of adsorption,
and strengthens the Au–C bond.
A similar effect has been found for adsorption of CO on small
anionic and cationic gold clusters71, 70.
It was found that the adsorption energy of CO
is larger in the case of a cluster of cations and smaller in the case of anions.
With the increasing cluster size, the differences in adsorption energies for
different charge states become smaller71.
It is important to mention that
strengthening the Au–C bond for C22+\nolinebreakH42+ adsorbed on the gold clusters
with an excess of the positive charge
is responsible for a cooperative effect in simultaneous adsorption
of C22+\nolinebreakH42+ and O22+ on neutral gold clusters.43
Indeed, the adsorption of O22+ on Aun2+
results in an
electron transfer from the gold cluster to the
anti-bonding 2π∗ orbital of O22+. In this case, C22+\nolinebreakH42+ effectively
adsorbs on the positively charged gold cluster.
A similar effect has also been predicted for adsorption of propene on O22+−\nolinebreakAun2+
clusters.40
It was supposed that the binding of propene to O22+−\nolinebreakAun2+ should be stronger
than the binding to the Aun2+ cluster due to the effective charge transfer from the
gold cluster to the oxygen molecule.40
The excess of the positive or the negative charge on gold clusters
results in a considerable change in the adsorption
energy of the di-σ-bonded C22+\nolinebreakH42+.
As has been discussed above, the adsorption energy of the
di-σ-bonded C22+\nolinebreakH42+ on small neutral gold clusters
exhibit an odd-even oscillations as a function of the cluster size.
These oscillations appear as a result of the electronic shell
effects in gold clusters: Aun2+ with odd n have one uncoupled electron
that can easily transfer to the empty π∗ anti-bonding orbital of C22+\nolinebreakH42+.
Such an electron transfer favors the di-σ configuration of the adsorbed C22+\nolinebreakH42+.
8a demonstrates that
there is no odd-even oscillations in the adsorption energy
of the di-σ-bonded C22+\nolinebreakH42+ on the anionic {{\text{Au}_{\vphantom{\text{}}\text{n}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}} clusters.
In that case, the cluster always possesses a weakly bounded electron that it can transfer to
the adsorbate.
In the case of the cationic Aun2++ clusters,
the electron back-donation process to C22+\nolinebreakH42+ is suppressed for any n.
8b shows that Eads calculated for
the di-σ-bonded C22+\nolinebreakH42+ on Aun2++ exhibit two profound
minima at n=3 and 9.
In accordance with the spherical jellium model,
Au32++ and Au92++ clusters possess closed shell
electronic structure with 2 and 8 valence electrons, respectively.
Hence, Au32++ and Au92++ are chemically inactive,
in particular in the processes where electron transfer from the cluster to the
adsorbate is responsible for formation of the chemical bond.
8c and 8d demonstrate
the n-evolution of the C–C bond distance, rC−C, in
C22+\nolinebreakH42+ adsorbed on {{\text{Au}_{\vphantom{\text{}}\text{n}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}} and Aun2++ clusters, respectively.
The C–C bond in the π-bonded C22+\nolinebreakH42+ on the anionic and the cationic clusters
is enlarged up to 1.36–1.40 Å irrespective to cluster charge.
This is 0.04 - 0.08 Å larger than that for the free C22+\nolinebreakH42+ (1.32 Å).
The di-σ-bonded ethylene is activated strongly in the case of adsorption on
the anionic {{\text{Au}_{\vphantom{\text{}}\text{n}}^{\vphantom{\text{}}\vphantom{\text{\smash[t]{2+}}}\text{\,,−−,}}}} clusters.
Thus, in the di-σ-bonded ethylene the C–C bond
is enlarged up to 1.53–1.42 Å for the anionic and only up to 1.48–1.37 Å for the
cationic gold clusters. Such an effect has a clear explanation.
The excess of electrons on a cluster promotes back-donation of an electron
from the cluster to the π∗ anti-bonding orbital of C22+\nolinebreakH42+, hence promoting its
activation. The excess of the positive charge on a cluster suppresses the back-donation process, thereby suppressing the C22+\nolinebreakH42+ activation.
Thus, by changing the cluster charge one can manipulate the cluster reactivity.
3.3 Vibrational spectra of ethylene adsorbed on small gold clusters
Analysis of the vibrational spectrum of adsorbed C22+\nolinebreakH42+ directly reveals
the mode of C22+\nolinebreakH42+ adsorption and the state of hybridization. This is due to the
sensitivity of the C–C stretching frequency to the hybridized state of carbon
atoms.11
In 1985 Stuve and Madix introduced the so-called πσ parameter
for characterizing rehybridization in C22+\nolinebreakH42+ upon adsorption
on metal surfaces.11
The πσ parameter depends on the
shift of the C–C stretching frequencies in C22+\nolinebreakH42+ and ranges from
0 for the free C22+\nolinebreakH42+, to 0.38 for K\nolinebreak[\nolinebreak(C22+\nolinebreakH42+\nolinebreak)\nolinebreakPtCl32+\nolinebreak] (Zeise’s salt),
and finally to 1 for C22+\nolinebreakH42+\nolinebreakBr22+.
Zeise’s salt and C22+\nolinebreakH42+\nolinebreakBr22+ have been chosen by Stuve and Madix as models describing
pure π- and di-σ-bonding of C22+\nolinebreakH42+, respectively.
Thus the larger the shift in vibrational frequencies
(compared to the free C22+\nolinebreakH42+), the greater the degree of rehybridization.
A similar approach can be applied for C22+\nolinebreakH42+ adsorbed on metal clusters.
1 presents the fundamental frequencies of free C22+\nolinebreakH42+ calculated in
the harmonic approximation and obtained from experiment72.
The obtained theoretical results are in a good agreement with experimental data.
Some discrepancy, especially noticeable
in the high frequency range, might result from anharmonic effects.
The anharmonic corrections can be taken into account, for example by the method
of n-mode coupling representations of the quadratic force field.73
However such a consideration goes far beyond the scope
of the present paper. In order to distinguish between π and di-σ modes of
C22+\nolinebreakH42+ adsorption one should to calculate the shift in the C–C stretching
frequencies of the adsorbed C22+\nolinebreakH42+ with respect to the free molecule. In that case the
anharmonic corrections to the vibrational energy levels can be partly canceled. Therefore in this
paper, we perform analysis of the vibrational spectra of the adsorbed C22+\nolinebreakH42+
in the harmonic approximation.
Following Ref.11, we have selected two vibrational modes:
ν6 and ν8, corresponding to the direct C–C and the in-plane CH22+
scissoring modes of vibrations, respectively.
Information on the shift of the vibrational frequencies of C22+\nolinebreakH42+ upon its adsorption
can be obtained from infrared or Raman spectroscopy.
9 presents the frequency dependence
of the Raman scattering activity calculated for a
C22+\nolinebreakH42+ molecule adsorbed in the π (9a) and di-σ (9b) configurations
on the neutral Aun2+ clusters with n = 1, …, 10.
The stretching modes corresponding to the ν6 and ν8 vibrations in C22+\nolinebreakH42+
are marked by the vertical arrows.
It is seen from 9a that the spectral dependences of the
Raman scattering activity calculated for the π-bonded ethylene on Aun2+ clusters
are similar for all the cluster sizes n considered.
The first maximum at 960 cm*-1* corresponds to the symmetric and asymmetric
swinging of H atoms in the direction perpendicular to the plane of C22+\nolinebreakH42+—
similar to the ν2 and ν3 vibrational modes in free C22+\nolinebreakH42+.
These vibrations remain unchanged in the π-bonded C22+\nolinebreakH42+.
Two intensive maxima at 1280 cm*-1* and 1560 cm*-1*
correspond to the ν6 and ν8 vibrational modes in C22+\nolinebreakH42+. These lines
are red-shifted in comparison with the free C22+\nolinebreakH42+.
Solid lines in 10 demonstrate shifts
Δν6π and Δν8π in harmonic frequencies ν6 and ν8
due to the π mode of C22+\nolinebreakH42+ adsorption on small neutral gold clusters.
For the π-bonded C22+\nolinebreakH42+, the scissoring mode ν8 is shifted by 30-40 cm*-1*
towards low frequencies if compared with the ν6 mode.
Both the Δν6π and Δν8π shifts
vary in the range of 58 - 142 cm*-1* and exhibit similar dependence
on the cluster size with local minima at n=6 and 8. The relatively small frequency shifts
Δν6π and Δν8π correspond to the values of the πσ
parameter typical for the π-bonded C22+\nolinebreakH42+.
9 demonstrates that the spectral behavior
of the Raman scattering activity calculated for
the di-σ-bonded C22+\nolinebreakH42+ is different from that obtained for
the π-bonded C22+\nolinebreakH42+.
Thus, for cluster sizes n= 2, 4, 5, 7 and 9,
a broad band appears in the frequency region of 900 - 1300 cm*-1*.
This band results from the strong mixing of the C–C stretching mode ν6
with the hydrogen swinging (ν2 and ν3) and twisting (ν4) modes.
The coupling of the ν2, ν3, ν4, and ν6 vibrational modes occurs
as a result of considerable deformations in C22+\nolinebreakH42+. These deformations include
the bending of H atoms in respect to the plane of a free C22+\nolinebreakH42+
due to the sp3 hybridization in di-σ bonded
C22+\nolinebreakH42+, as well as the twisting of CH22+ groups in respect to each other along the C–C bond.
Such twisting is
especially noticeable for C22+\nolinebreakH42+−\nolinebreakAu22+ and C22+\nolinebreakH42+−\nolinebreakAu42+ clusters.
The strong coupling between different vibrational modes makes identification
of ν6 and ν8 vibrations to be rather difficult: the mixed modes
form new levels, neither of which represent the true ν6 and ν8 states.
Appearance of the sp3 hybridization character in di-σ bonded
C22+\nolinebreakH42+ results in a considerable red-shift in the stretching frequencies.
Such shifts Δν6di−σ and Δν8di−σ
in harmonic frequencies ν6 and ν8 are shown by the dashed lines in 10.
It is seen that the Δν6di−σ shift is considerably larger than the
Δν8di−σ which can be explained by the strong coupling between the
ν2, ν3, ν4, and ν6 vibrational levels.
The Δν6di−σ and Δν8di−σ frequency
shifts varies in the range of 200 - 370 cm*-1* for n=2, … ,10.
These values correspond to the πσ
parameter typical for the di-σ-bonded C22+\nolinebreakH42+. Thus,
the analysis of the vibrational modes of C22+\nolinebreakH42+ adsorbed on gold clusters
allows one to identify the specific modes of C22+\nolinebreakH42+ adsorption and the
rate of rehybridization upon adsorption.
4 Conclusions
In summary, we have demonstrated that C22+\nolinebreakH42+ can be adsorbed
on small gold clusters in two different configurations,
corresponding to the π- and di-σ-bonded species.
Adsorption in the π-bonded mode dominates over the di-σ
mode over all considered cluster sizes n, with the exception of the
neutral C22+\nolinebreakH42+−\nolinebreakAu52+ system. We found a striking difference in
the size dependence of the
adsorption energy of C22+\nolinebreakH42+ bonded to neutral gold clusters in the
π and di-σ configurations. The adsorption energy, calculated
for the di-σ-bonded C22+\nolinebreakH42+ exhibits
pronounced odd-even oscillations, showing the importance of the electronic
shell effects in the di-σ mode of ethylene adsorption on gold clusters.
The different rate of hybridization in the π- and di-σ-bonded
C22+\nolinebreakH42+ can be responsible for the different rate of catalytic activation
and hence, different reactivity of adsorbed C22+\nolinebreakH42+.
We have also demonstrated that the interaction of C22+\nolinebreakH42+ with small gold clusters
strongly depends on the cluster charge. Hence ethylene adsorption and reactivity
can be manipulated by the cluster charge.
The strengthening of the ethylene – cluster interaction for gold clusters
with an excess of the positive charge is responsible for the cooperative effect
in simultaneous adsorption of C22+\nolinebreakH42+ and O22+ on neutral gold clusters.
This effect can play an important role in the mechanism of
catalytic oxidation of alkenes by dioxygen on the surface of gold
clusters.
Finally we have shown that the analysis of the vibrational modes of C22+\nolinebreakH42+
adsorbed on gold clusters allows one to identify the specific modes
of C22+\nolinebreakH42+ adsorption, the rate of rehybridization upon adsorption, and thus the
C22+\nolinebreakH42+ reactivity.
In the present work, we have considered adsorption of ethylene molecule
onto small gold clusters consisting of up to 10 atoms.
Many interesting problems beyond the scope
of the present work arise when considering multiple molecular
adsorption on gold nanoparticles of larger
sizes up to 1-5 nm, where the strong dependence of the catalytic
activity of gold nanoparticles has been observed experimentally.
In particular, multiple molecular adsorption on the surface of
gold nanoparticles can result in a considerable change in
cluster morphology, even for relatively large cluster sizes.
Another important direction for future development is to
investigate the effect of alloying on the adsorption and catalytic activation
of alkenes on gold clusters. The doping of a gold cluster with atomic impurity
can result in considerable changes in the geometrical and electronic structure of the cluster,
thereby modifying and controlling the catalytic activity.
Understanding how to enhance and control chemical reactions on a cluster surface
is therefore a vital task for nanocatalysis.
{acknowledgement}
This work was supported by the Global COE Program
(Project No. B01: Catalysis as the Basis for Innovation
in Materials Science) from the Ministry of Education,
Culture, Sports, Science and Technology, Japan.
Calculations were performed in the Research Center for Computational Sciences,
Okazaki Research Facilities.

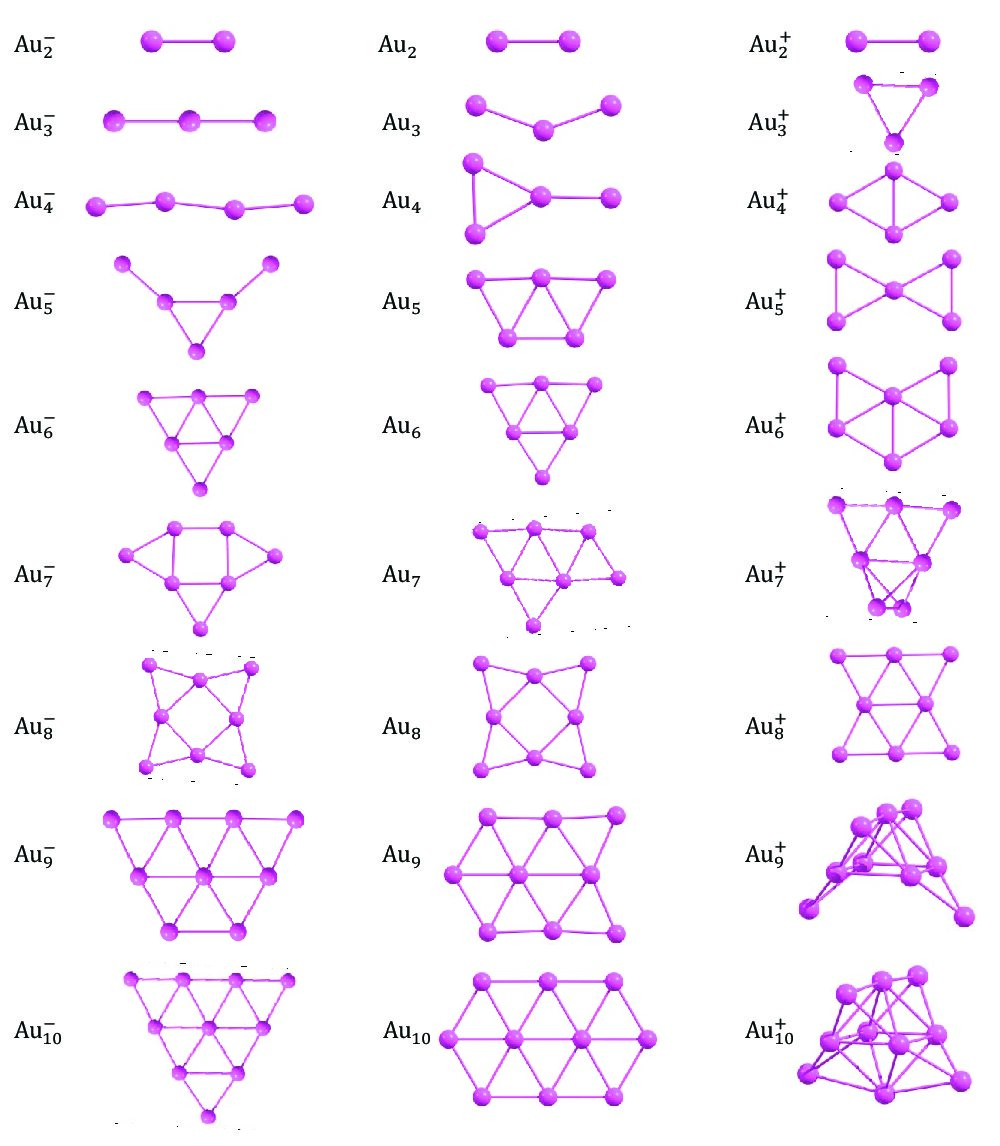 Figure 0
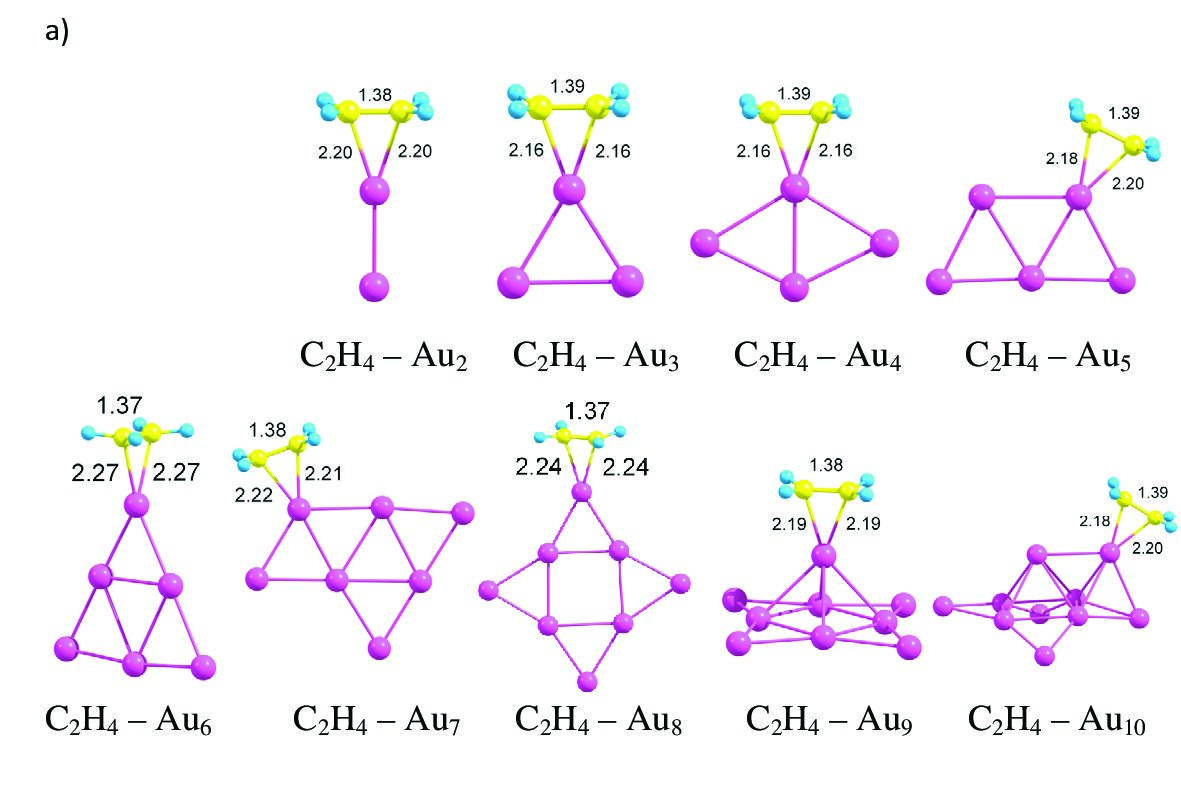 Figure 1
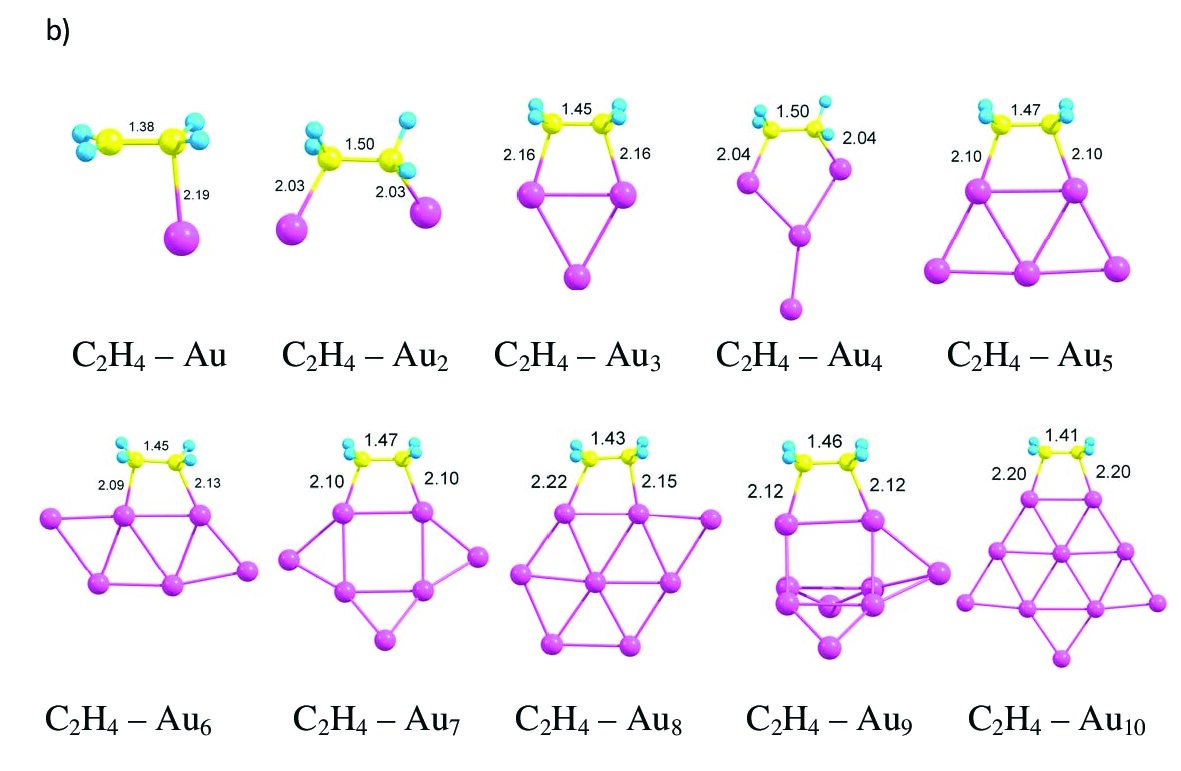 Figure 1
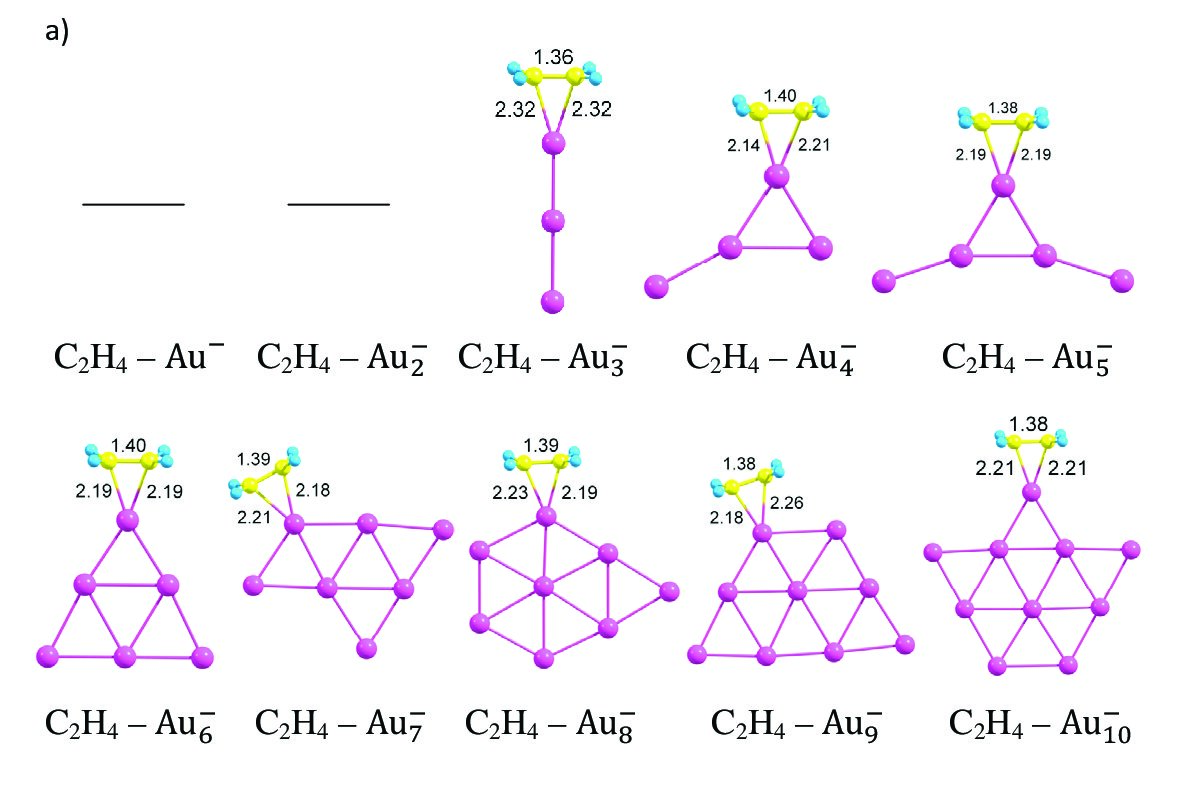 Figure 2
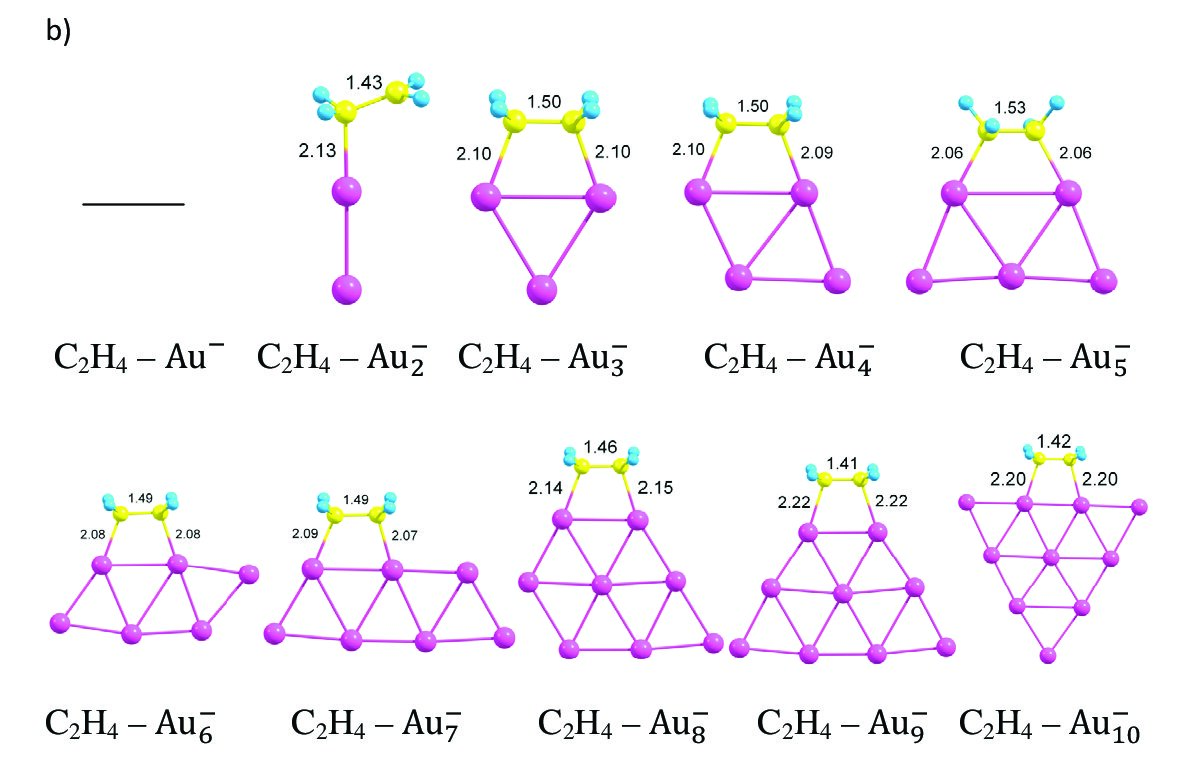 Figure 2
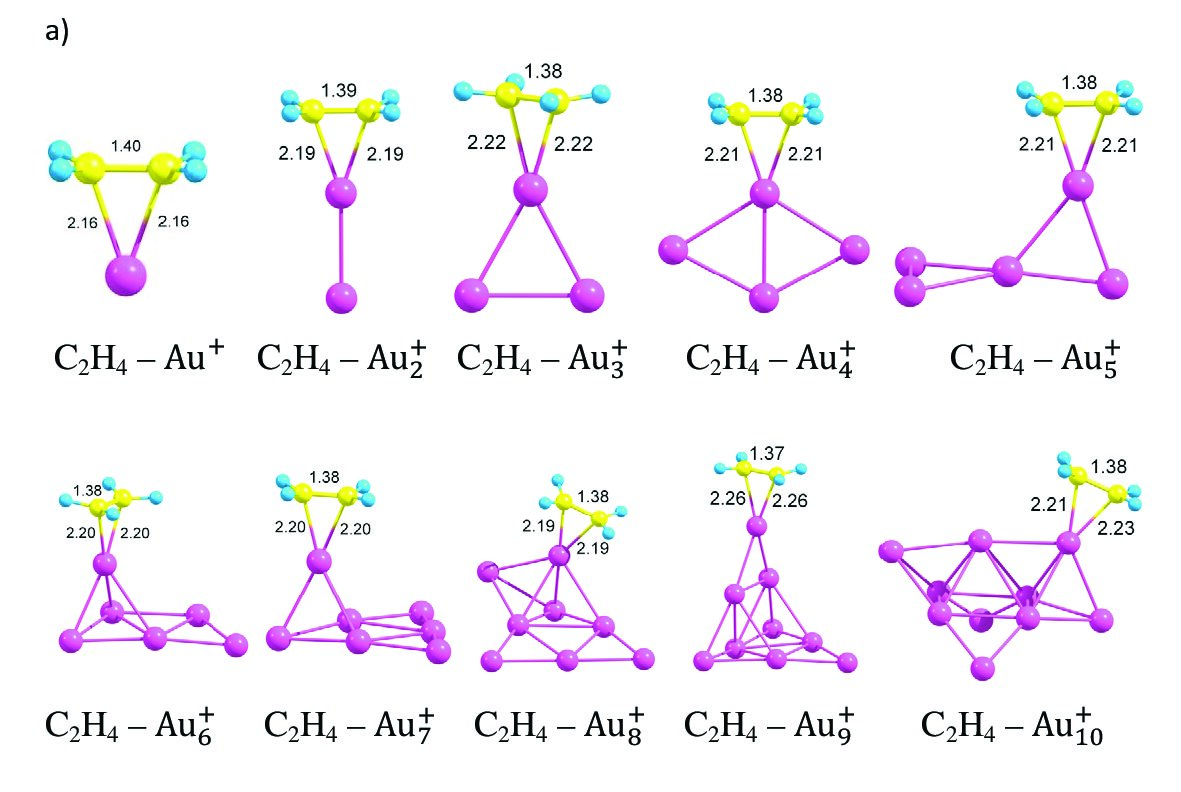 Figure 3
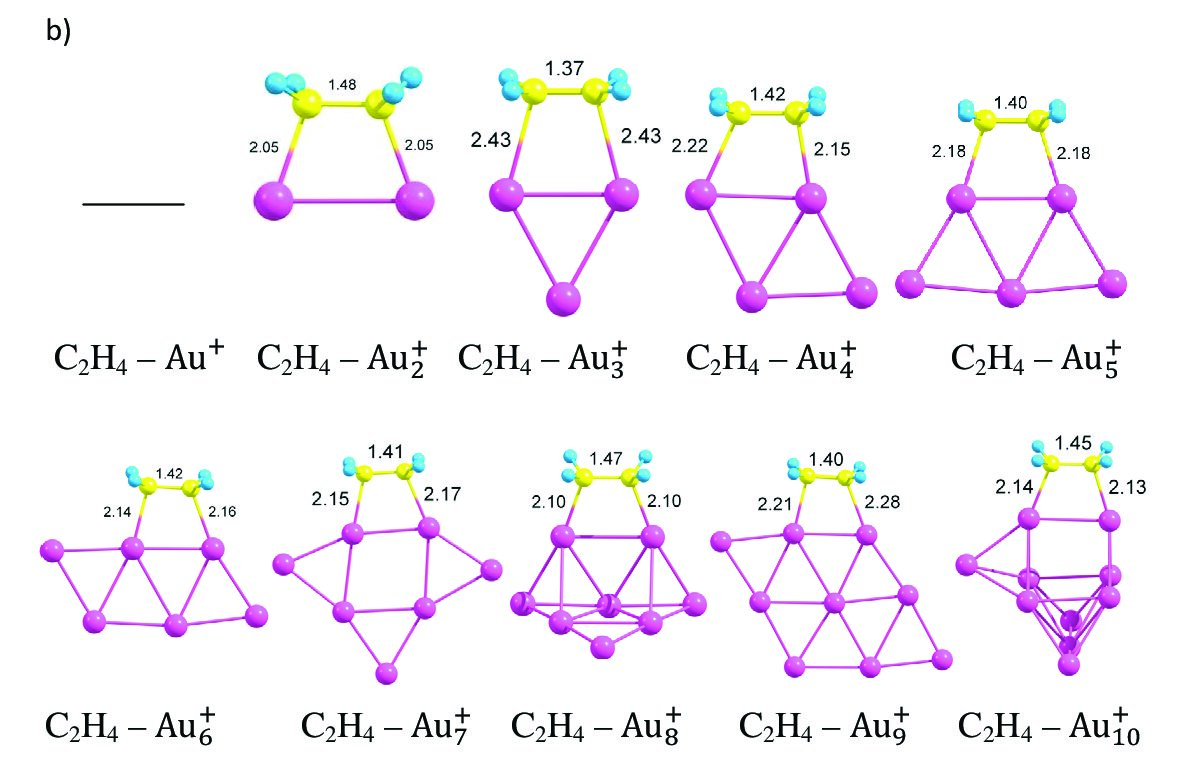 Figure 3
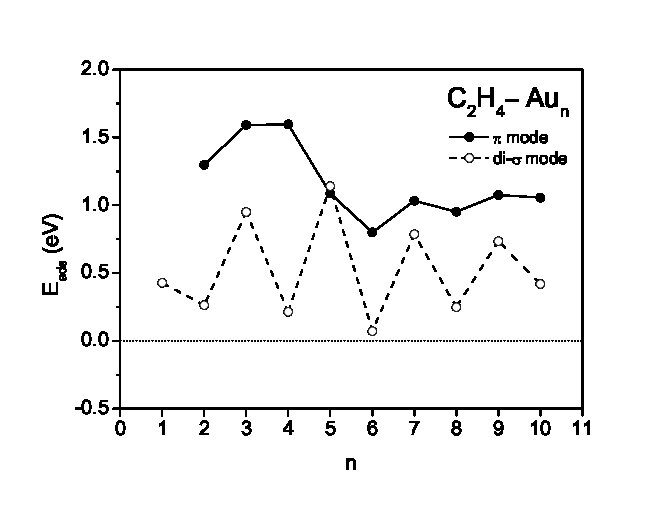 Figure 4
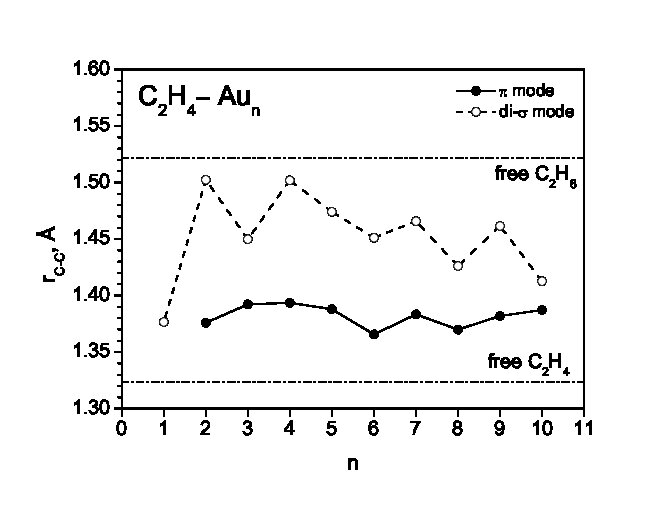 Figure 5
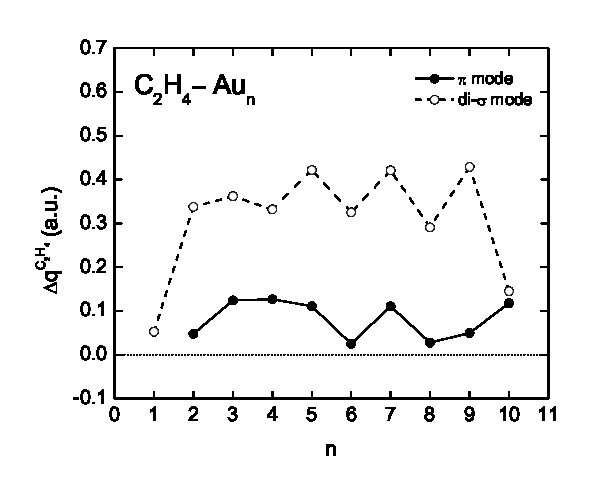 Figure 6
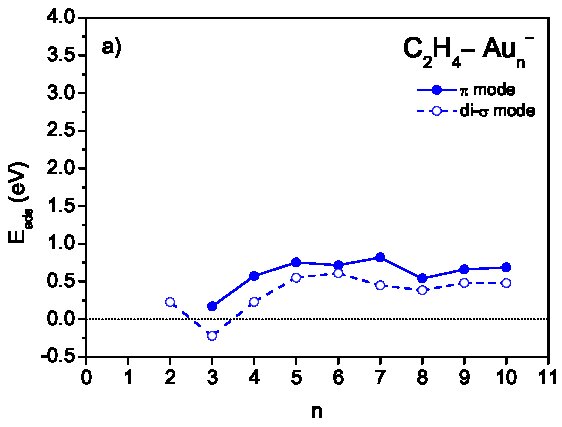 Figure 7
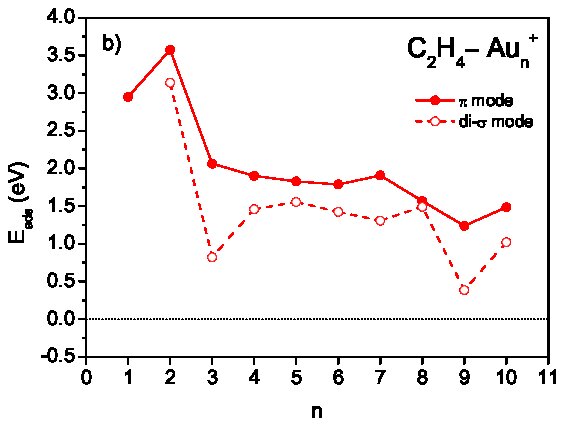 Figure 7
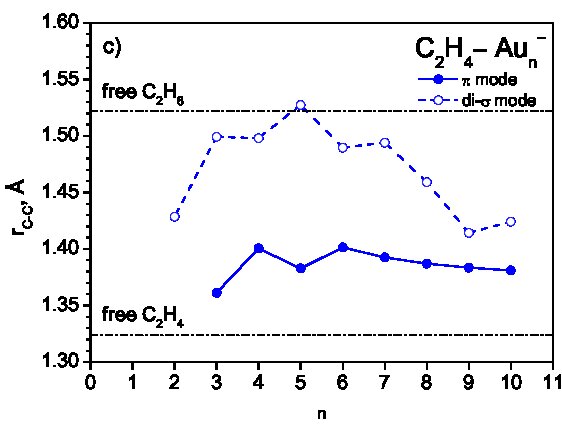 Figure 7
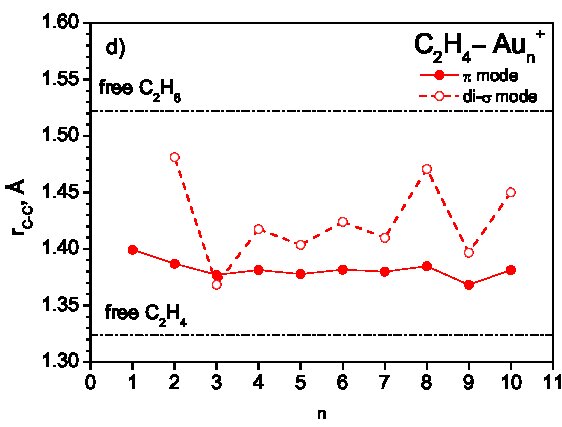 Figure 7
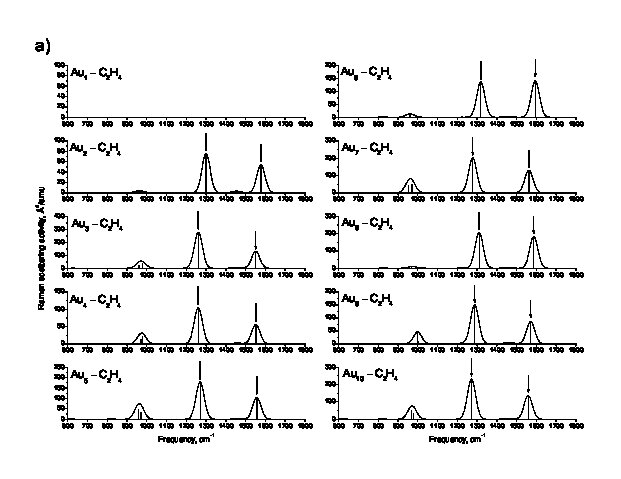 Figure 8
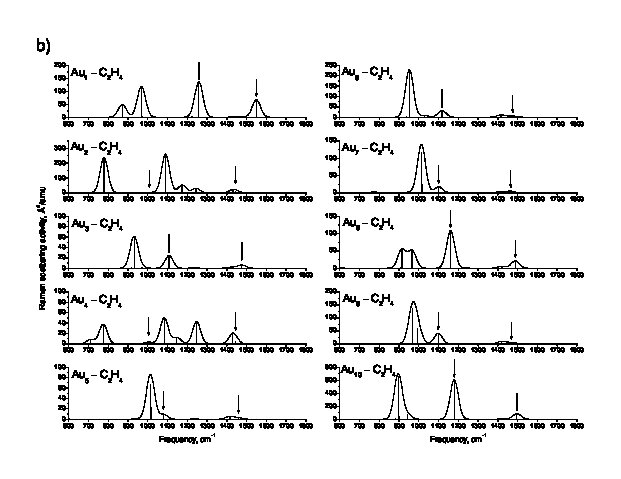 Figure 8
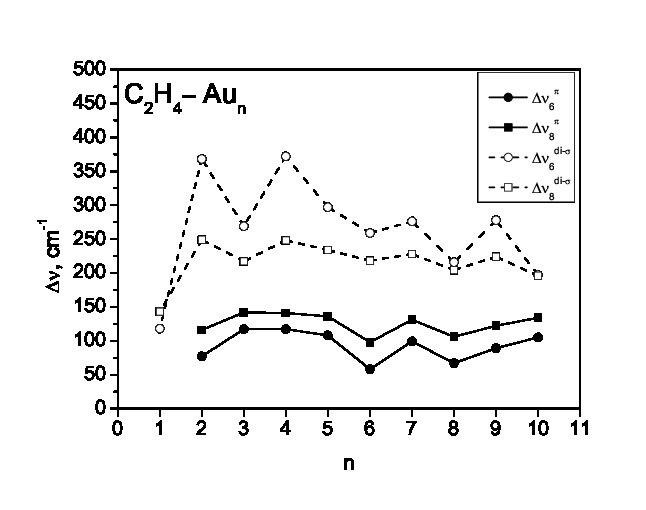 Figure 9